
GMP, Good Manufacturing Practice, is the rulebook the FDA, EMA, and pretty much every regulator on earth uses to decide whether a pharmaceutical product can be sold. The point isn't that the medicine works. Clinical trials prove that. GMP proves the medicine you ship is the same medicine you tested, every time, every batch, every year.
Want one partner for design, components and packing? See pharmaceutical contract packaging at Ecobliss.
It sounds bureaucratic. It is. But the stakes are simple: a single GMP failure in raw materials, equipment cleaning, or batch documentation can pull a product off the market, trigger a recall, or block an approval that took ten years and a billion dollars to earn.
Where GMP actually lives in the law
In the US, in 21 CFR Parts 210 and 211, written and enforced by the FDA. In the EU, in EudraLex Volume 4, written by the Commission and enforced by national competent authorities. In Japan, in PMD Act and PIC/S guidelines. The texts are different but the principles overlap so heavily that a facility certified to one usually qualifies for the others with minor adjustments.
How 21 CFR Parts 210 and 211 are organized
Part 210 sets the minimum current GMP for manufacturing, processing, packing or holding drugs: every drug product has to be made under conditions that ensure its identity, strength, quality and purity. Part 211 fills in the specifics, subpart by subpart:
- B, Organization and personnel: a quality unit, independent of production, with authority to release or reject batches, plus training for everyone who works in a regulated process.
- C, Buildings and facilities: facility design, cross-contamination prevention and environmental controls.
- D, Equipment: qualification (IQ, OQ, PQ), cleaning and calibration.
- E, Control of components: handling, sampling, testing and release of raw materials, the chain that lets you reconstruct any batch.
- F, Production and process controls: batch records, in-process testing and written procedures.
- G, Packaging and labeling: packaging operations, label control and finished product handling.
- H, Holding and distribution: warehouse conditions, distribution records and quarantine.
- I, Laboratory controls: test methods, stability programs and retention samples.
- J, Records and reports: how long records are kept and how they stay accessible.
What GMP actually requires
It boils down to: prove that what you say you did, you did. Every facility, every piece of equipment, every batch, every employee, every deviation, every cleaning cycle. Documented, verifiable, reconstructable.
Specifically:
- A quality unit independent of production that can refuse to release a batch.
- Validated processes, meaning documented evidence the process consistently produces what you claim.
- Qualified equipment, IQ-OQ-PQ run on every machine that touches a regulated product.
- Trained staff, with retraining records when procedures change.
- Cleanroom controls, with environmental monitoring graded by ISO 14644 class.
- Batch records that survive an FDA 483 inspection a decade later.
What GMP looks like in practice
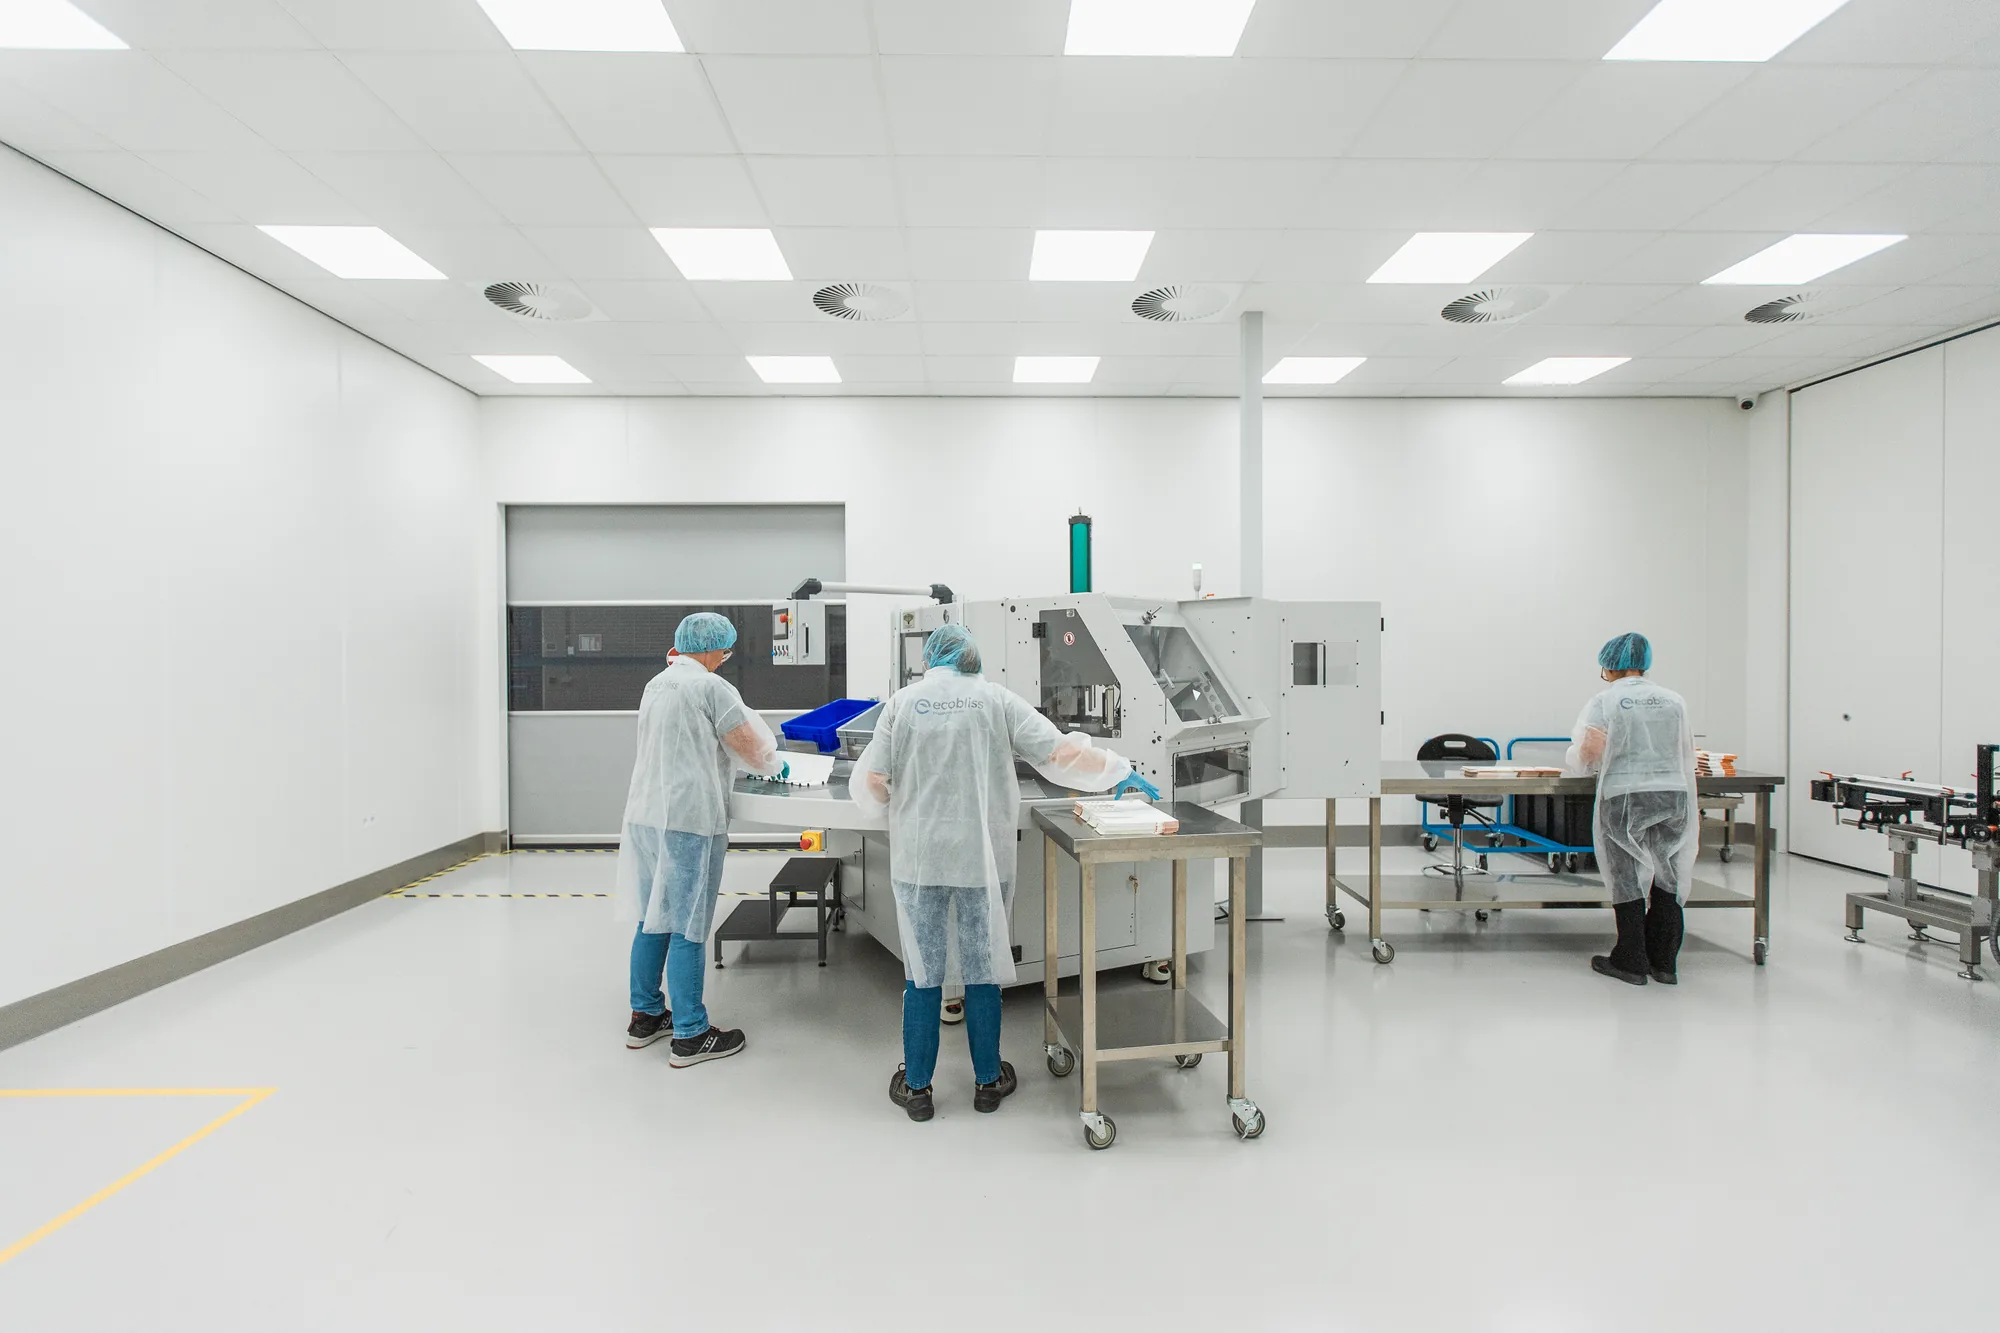
A morning on a GMP packaging line: an operator scans her badge, the line software checks her current GMP training certificate, she's allowed to start. The cold seal machine boots and runs a self-check. The first ten wallets out of the line go to the QC operator who measures seal integrity, prints, and labels them. Each wallet has a 2D Data Matrix code, registered against the batch number, registered against the API lot, registered against the supplier.
Three thousand wallets later, the line stops, the operator records why, the deviation goes into the batch record. By the end of the shift the batch record is sixty pages, signed off by quality, sealed against tampering. If a recall happens in 2031, that file gets pulled and reviewed.
What an FDA inspector actually looks at
Inspectors do not read all of Part 211 against your facility; they sample. The findings that show up most often in FDA 483 observations:
- Batch records that do not support reconstruction of the batch, with missing signatures or incomplete deviation entries.
- Cleaning validation gaps: no documented, current evidence that cleaning removes residues to acceptable limits.
- Training records that are out of date for operators on the line.
- Deviations that are not escalated, investigated or closed within a defined timeframe.
- Environmental monitoring data that does not show consistent control, or trends that were ignored.
What GMP means for the packaging side
Packaging is where most GMP failures show up in audits, because it's the layer that touches the lot number, the patient information, the serialization code, and the tamper seal. A misprinted leaflet, a missing 2D code, a mixed-up batch label, all of those are GMP findings even if the API itself is perfect.
Label artwork changes that skipped change control are another common packaging finding. All of these are preventable.
If you're picking a contract packer, the question isn't "are you GMP-licensed." Everyone says yes. The real questions are: when was your last FDA inspection, how many 483 observations did you take, and can you walk me through your last deviation. If you want our answers, we'll send you the inspection record.